
Overview.
Post-excitation dynamics in biomimetic photo-catalysts. Strain-induced metal insulator transitions in strongly correlated transition metal oxide materials. Tracking ultrafast charge migration at a molecule-material interface with x-ray spectroscopy. Due in large part to the number of and complicated interactions between the electrons in systems like these, current theoretical methods are often unable to offer reliable predictions in support of the design and analysis of new experiments and technology. Our group specializes in developing new theoretical models, algorithms, and software in order to achieve reliable predictive power over ever larger and more complicated collections of electrons.
Treating Excited States on an Equal Footing.
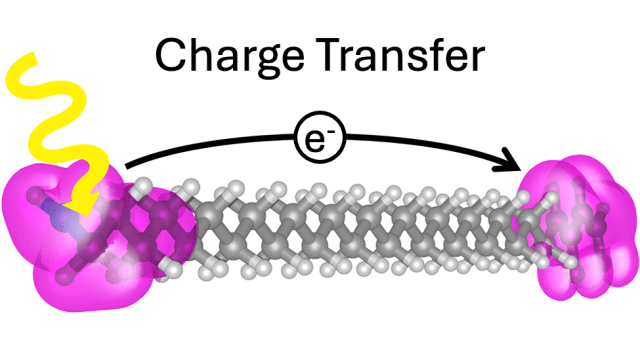
Due to both the historical priority of ground states and the inherent limitations of the traditional variational principle, electronic structure methods for modeling excited states tend to be more approximate than those for ground states. This imbalance is present both in chemistry, where excitations are most often modeled by linear response theory, and in solid state physics, where band gaps and spectra are most commonly derived from DFT and the many-body perturbation theory of Greens functions. While substantially more robust methods exist in both fields for modeling ground states (e.g. DMRG, QMC, fully relaxed coupled cluster, and so on), applying these methods to excited states is typically not possible without additional approximations, such as the linear response approximation or the use of state-averaged orbital shapes. A major direction of investigation in our group is into methods that directly target excited states through modifications to the ansatz, thus avoiding the need for different levels of approximation for different states while maintaining accuracy comparable to that of the ground state. These approaches are especially important for excitations that frustrate traditional approaches, such as charge transfer excitations, band gaps near a Mott transition, and double excitations.
Exploiting the Locality of Reactions.
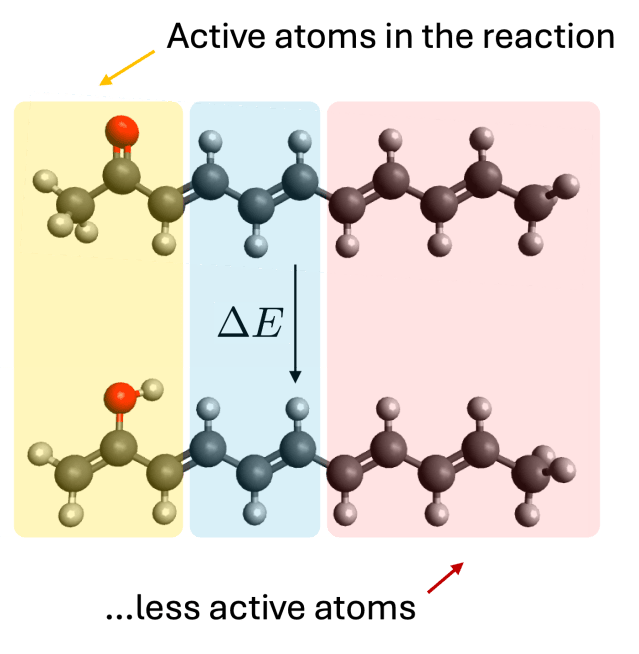
In chemistry, one is rarely interested in the energy needed to fully pull apart a molecule into well separated electrons and nuclei (what many practitioners call the absolute energy), but instead in specific energy differences that arise from rearranging bonds or absorbing photons. However, to calculate these differences, the most common approach is to compute absolute energies of the complete systems both before and after the desired reaction, even though in most scenarios it is only a small portion of the molecule that actually undergoes significant change. We are increasingly interested in exploring approximations that tailored to predicting accurate energy differences while intentionally neglecting wave function details that are important for total energies but are expected to cancel out when the reaction or excitation energy is evaluated. The relative locality of many chemical processes offers us a strong lever towards these ends, and we are having fun getting confused about how to use that lever aggressively but responsibly to make unusually efficient predictions whose absolute energies are terrible but whose relative energies are accurate.
Handling the Breakdown of Single Configurational Character.
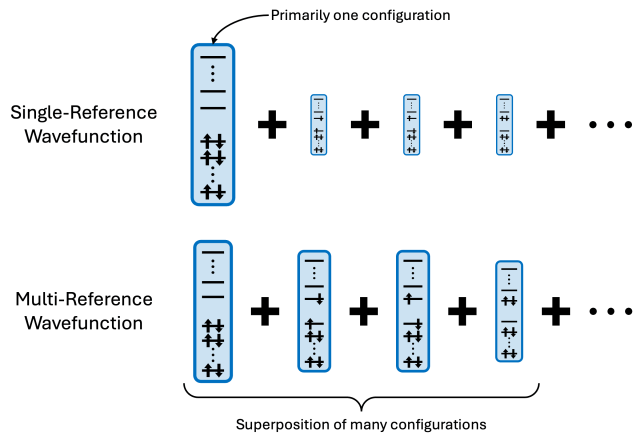
A significant portion of organic chemistry can be handled by ground state methods that assume a single configuration of the electrons in the MOs is a reasonably accurate picture. If one were only ever interested in the stable conformations of the reactants or products, this assumption is usually safe. However, if we instead wanted to map out the bond breaking and forming procedure, multiple electronic configurations are likely to make large contributions to the wave function. Such multi-reference character is also present in many inorganic systems involving transition metal complexes, where the locality and narrow angular distributions of d and f electrons often prevent the large HOMO/LUMO splittings found in stable organic molecules. Although this situation offers a rich environment for developing catalysts and other technologies, it plays havoc with standard electronic structure methods like DFT. We continue to be curious about how to make progress in this area of modeling, especially in the context of photochemistry and the challenge of capturing both weak and strong correlation effects simultaneously.
 Neuscamman Group
